


The author, evil genius, has said that you're saving your life at a low point in your life, but you've been at a low point, and people can't stand it. Today we begin the second step of molecular dynamics: defining unit boxes and adding solvents. The last thing we did was create protein popping structures, articles. Gmx pdb2gmx reads. Pdb or. Gro files and reads database files, adding hydrogen to molecules and generating coordinates in gromacs (glomos) or optional. Pdb, format and gromacs scaling. You can then process these files to generate running input files. On that basis, then, we have to move on. The first output log needs information
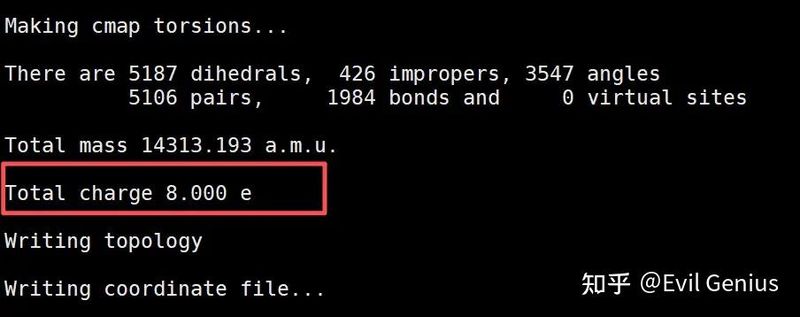
Then we'll have to balance the charge. Let's start with the argument gmx editconf to convert the common structure format to. Gro, g96 or. Pdb in molecular dynamics simulations, a cyclical simulation box is usually added to the system. Gmx editconf has many control box options.
Options to feature input files:
[ chuckles ]<.gro>(conf. Gro)
structure file: gro g96 pdb
-n-n<.ndx>(index. Ndx) (opt.)
index file
[ chuckles ]<.dat>(bfact. Dat) (opt.)
general data file
options to specify output files:
- oh, no<.gro>(out. Gro)
structure file: gra g96 pdb
- mead<.pqr>(mead. Pqr)
cordonate file for mead
other options:
-[no]w (no)
vew output, xvg, xpm, . Pdb files
-[no]ndef (no)
close output from default index groups
-bt(triclinic)
box type for-box and-d: technical, cubic, dodecahedron,
octahedron
-box(0)
box vector lengths (a,b,c)
- angles(90 90 90)
angles between the box vectors (bc, ac, ab)
- d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-d-(0)
distinction between the system and the box
-[no]c (no)
center molecule in box
- center(0)
shift the global center to (x, y, z)
-aligncenter(0)
center of roation for identity
-align(0)
allan to target vector
- translate(0)
transport
- lotate(0)
rotation around the x, y and z axes in damages
-[no]princ (no)
orient molecule (s) along their principal axes
- scale(1 1)
scaling factor
- density(1,000)
density (g/l) of the output box achieved by scaling
- [no] pbc (no)
remove the human whole again
- yes, sir(-1)
remember reidous starting from renr
-[no]grass (no)
store the charge of the atom in the b-factor field and the radius
of the atom in the occupancy field
- rvdw(0. 12)
default van der waals radius (in nm) if one can not be found in the
databIf no parents are present in the global file
-[no] sig56 (no)
use rmin/2 (minimum in the van der waals power)
sigma/2
-[no]vdwread (no)
read the van der waals radio from the file vdwradii. It's not like i'm going to have to do thisLet's go.
-[no]atom (no)
force b-factor attachment per atom
-[no]legend (no)
make b-factor legend
- label(a)
addlain label for all rezues
- [no]coNect (no)
add coNect records to a. Pdb file when writtenNo, i'm not. When a topology (tpr file) is present
known issues
? For complex problems, the physical removal route may break down ?
? In that case you can use gmx trjconv.
-f:
... Gro/. G96/...>
(conf. Gro) enter protein structure - o:
. Grol. G96/...>
(out. Gro) output structural file with analogue box information - c: place the protein molecule at the centre of the box - bt: box type, default use of long box, dodecahedron, cubic etc. - d: the distance between the solution and the box, the minimum distance in x, y, z direction to the edge of the box is an important parameter, as we use cyclical boundary conditions and must meet the minimum image agreement, i. E. A protein can never “see” its own cyclical image (which cannot interact with itself), otherwise the calculated strength will contain a false part ... The designated distance between the solvent and the box is 1. 0 nm, meaning that the distance between any two cyclical images of the protein molecule is at least 2. 0 nm. This distance is sufficient for any cut-off formula commonly used in the simulation. -d option sets the minimum distance from the molecule to the edge of the box, in nm, which determines the size of the box. In theory, in most systems, -d cannot be less than 0. 9 nm. When you learn more about cyclical boundary conditions and the type of box, you strongly recommend the use of a sorghum dodecahedron crystal because, in the case of cyclical distances, it is only 71 per cent of a cube crystal, thus reducing the number of solvent water molecules that need to be added. Let's expand on the cyclical boundary
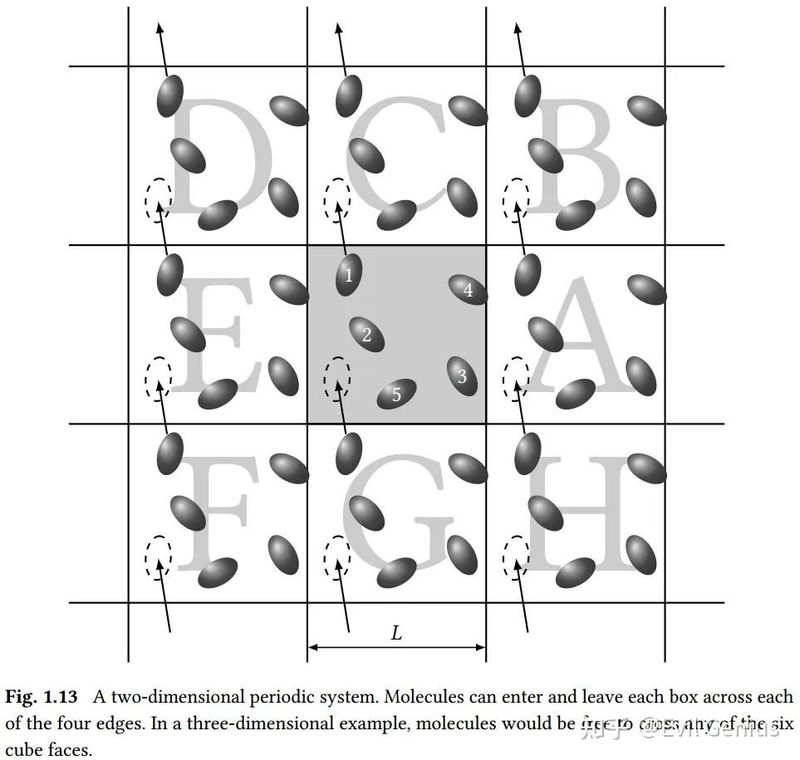
Considering a very small cube droplet unit, which contains a thousand atoms, 83 = 512 inside, which can be considered to be completely liquid, and the rest, almost half of it, will obviously not have the same properties on the skin of this unit. Even at a scale of 106 atoms, almost 6% have different properties. And what do we do with this liquid unit, which we have to do with a sample of macro-generality? The problem can be overcome by the cyclical boundary conditions proposed by born and von karman in 1912. The first is that, assuming that there are countless identical liquid cells in space, particles in the cube boundary can still be activated by nearby cells. The second is that there is still a way for us to get what the nearby cells are called mirrors of the body.
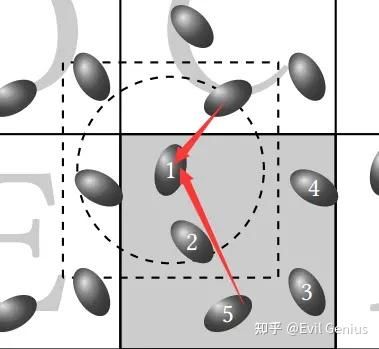
A very critical question is, can this very small, infinity cycle be of the same nature as the macro system? This depends on the range of power between molecules and the range studied. In general, for short-range strong interactions, the power distance cannot exceed half the box. This may be a little abstract, but look at fig1. 13, if the particle 1 is more than twice the size of the box, then the particle itself will work, obviously make sense; if more than half the box is long, then the particle 1 in the positive direction will work for it once, and again from the opposite direction, make no science sense.
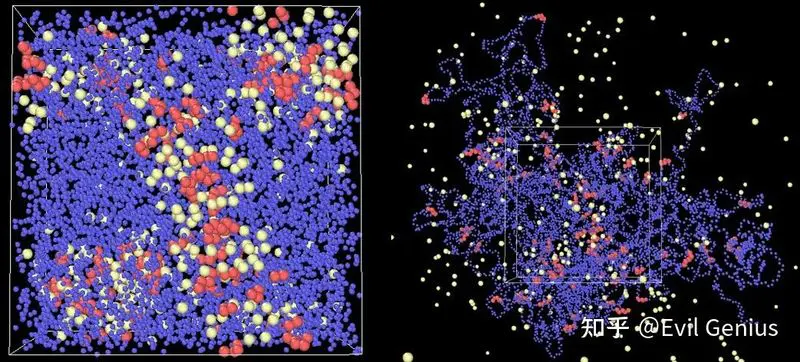
In the case of cubes, for example, we need to introduce the concept of image flag. This value represents the number of times the atom goes through the box, which is going through the image flag+1, the reverse direction-1. When the particle runs out of the box in the right direction, that means that it runs into the next box, the value of which is greater than l., which should be subtracted from the value of l, and where it actually goes, and the data can be used to calculate parameters such as proliferation
I'm not sure if you're going to be able to do thatThen we add the solvent gmx solvate, and we look at the parameters
Options to feature input files:
-cp<.gro>(protein. Gro)
structure file: gro g96 pdb
-cs<.gro>(spc216. Gro) (lib.)
structure file: gro g96 pdb
options to feature input/output files:
- p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-p-s-p-s-p-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-s-<.top>(topol. Top.)
topology file
options to specify output files:
- oh, no<.gro>(out. Gro)
structure file: gra g96 pdb
other options:
-box(0)
box size (in nm)
- radius(0. 105)
you're not going to get away with this.
- scale(0. 57)
scale factor to mostly van de waals radio from the databLet's go. The default value of 0. 57 yields
you're gonna have to do it.
- shell(0)
tickness of optioI'm sorry, my dear, but i'm sorry, but i'm sorry.
-maxsol(0)
maximum number of slovakians to add if they fit in the box. If zero (default) this is signed
-[no]vel (no)
keep plans from input solute and solvent
Gmx solvate can do one of the following two things: 1. Create a solvent-filled box. ... Can be done by specifying -cs and -box options. ... For structure documents with box information but not atoms can be accomplished by designating -cs and -cp. 2. Solve solvents of solvent molecules, such as proteins, into the encirclement of solvent molecules. -cp and -cs, respectively, are used to specify solubility and solvents. Without setting -box, the box information in the solvent coordinates file (-cp). -cp (protein. Gro) enters documents. The protein configuration document used is from the output document in the previous editconf step -cs (spc216. Gro) input library documents, and the formulation file of gromacs, which is standard installed -ppol. Top, specifies that the spoo-o (out ofgro) export the agent's structure file is a genericly balanced p21. C. Attention to topol. Top
I'm sorry
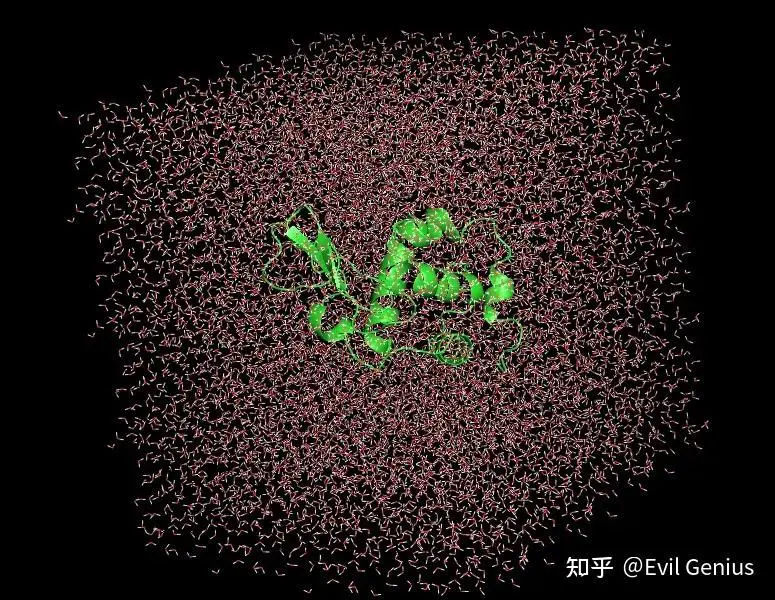
Solvate records the increase in the number of water molecules and puts them in the piston file to show the changes it has made, noting that if you use other (non-water) solvents, solvate does not write in the piston file! It automatically records that the function of updating water molecules is directly in the source code. Once the simulation box is defined, the solvent is added using the solvate module, and the structure file containing the solvent is exported solv. Gro.
Gmx solvate-cp newboxLook at the added effect
Life is good, with you better。









